Solid Phase Peptide Synthesis Impurity Profiles: Why the Impurities Are Predictable
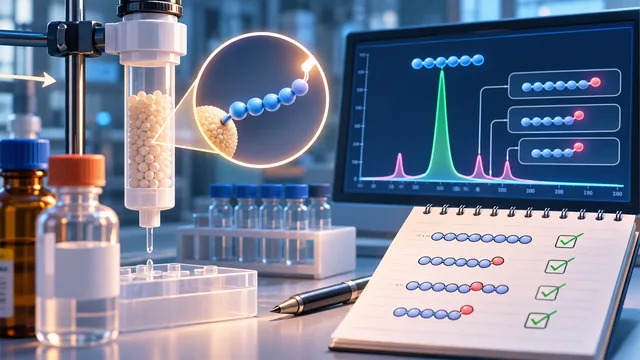
Short answer: the impurities in a synthetic peptide are a readable consequence of how it was made. Deletions come from incomplete coupling, truncations from chain termination, adducts from incomplete deprotection, and each type behaves differently in an assay. The profile is more informative than the headline purity figure.
- Solid phase synthesis builds the chain one residue at a time, and each cycle has a yield below one hundred percent.
- Deletion sequences differ from the target by one residue and can retain partial activity.
- Truncated sequences are usually inert, which makes them a dilution rather than a confound.
- Oxidation, racemization and adducts each leave a characteristic mass signature.
How does the synthesis create the impurities?
Solid phase synthesis anchors the first residue to a solid support and adds the rest sequentially. Each addition is a cycle: deprotect the growing chain's reactive end, couple the next protected residue, wash, repeat. At the end, the chain is cleaved from the support, the side-chain protecting groups are removed, and the crude material is purified.
Every cycle is a chemical reaction with a yield below one hundred percent, and the shortfalls accumulate. A small per-cycle loss across a long sequence produces a crude mixture in which the target is a minority species before purification. That is normal chemistry, not poor practice, and it is why purification is the expensive part.
What are the main impurity types and why do they matter differently?
Deletion sequences arise when a coupling step fails on some fraction of chains and synthesis continues, producing a product missing one internal residue. These are the impurities that matter most, because a sequence differing by one residue can retain measurable binding or activity, and it co-elutes closely with the target. A deletion impurity produces a plausible but shifted result rather than an obvious failure.
Truncated sequences arise from chain termination, leaving shorter fragments. These are usually biologically inert, so their practical effect is to make the vial less concentrated in active material than the nominal figure suggests.
Adducts arise from protecting groups that were not fully removed during cleavage, and they appear at predictable mass offsets. Oxidation products affect methionine, cysteine and tryptophan and typically present as an addition of sixteen mass units per oxygen. Racemization at susceptible positions produces diastereomers that are chemically identical in mass and may only separate under carefully chosen chromatographic conditions, which means a method that does not look for them will not find them.
Why does sequence length and composition change the risk?
Because difficulty is sequence-dependent. Long sequences accumulate more cycles and therefore more opportunity for deletion. Sequences with a propensity to form secondary structure on the resin couple poorly, because the growing chain aggregates and the reactive end becomes less accessible. Certain residue pairings are known to be difficult, and some side chains are more prone to side reactions during cleavage.
A laboratory can therefore anticipate the risk from the sequence alone, which is one reason the full sequence belongs on product documentation rather than a product name and a percentage. Where a supplier publishes the sequence with the lot analytical package, as Bluum Peptides does per product, the impurity profile can be interpreted rather than guessed at.
Can the profile be read off a certificate?
Partially, and more so when the underlying data is available. A purity percentage states how much of the eluted ultraviolet-absorbing material was the main peak. The chromatogram shows how many other peaks there are, how large, and crucially how close to the main peak, which is what indicates whether closely related species are present. Mass spectrometric data on the significant impurities identifies which type each one is.
That is why suppliers who provide the chromatogram and spectrum on request, rather than only a summary figure, enable a different quality of purchasing decision. A buyer can see whether the method was capable of resolving the impurities that matter for that sequence, instead of trusting that it was.
What should a laboratory do about it?
Write the specification as a set rather than a threshold: purity with method conditions, peptide content, identity confirmation, water content, and where it matters a limit on any single impurity rather than only on the total. Then qualify each new lot in the assay against the outgoing one, because two lots meeting the same total purity specification can carry different profiles and behave differently.
Frequently asked questions
Why is a longer peptide usually less pure?
Because each synthesis cycle has a yield below one hundred percent and a longer chain requires more cycles, so deletion and truncation species accumulate before purification.
Which impurity is most likely to affect an assay?
A deletion sequence, because it differs from the target by a single residue, often retains some activity, and co-elutes closely enough that a routine method may not resolve it.
Does high purity mean the impurity profile does not matter?
Not always. A small proportion of an active deletion species can matter more in a sensitive assay than a larger proportion of inert truncated fragments, which is why the profile is more informative than the total.
Can racemization be detected by standard chromatography?
Only with conditions chosen to resolve diastereomers. Mass spectrometry cannot distinguish them, since the mass is unchanged, so a method that was not designed to look for racemization will report none.
This article is provided for research and informational purposes only. The materials discussed are laboratory reagents intended for in vitro and preclinical research use. Nothing here describes or endorses use in humans, and no claim is made regarding any outcome, benefit, or application beyond laboratory research.